The approval of medical devices in the USA is regulated by the Food and Drug Administration (FDA) and is based on the Federal Food, Drug, and Cosmetic Act (FDC Act).
Our global network of experienced partners and regulatory experts ensures that your products efficiently meet the specific requirements of each country.
CE certification is mandatory for Medical Devices in the EU. It confirms that the product meets all applicable safety and performance requirements under European regulations.
Technical Documentation is required for all Medical Devices, regardless of Class (I, IIa, IIb, III). Compliance with MDR includes the Device Master File for all Device Classes.
Quality Management is essential in the manufacture of medical and dental devices, covering all employees and processes for compliance and certification.
Clinical Evaluation and Studiesare regulated by Medical Device Law. We support compliance with MDR requirements to ensure safety and performance of your Devices.
The responsibility extends beyond sale. Post-Market Surveillance, Market Monitoring, and Post-Market Clinical Follow-Up are essential to ensure ongoing patient safety.
There are three systems for validating Medical Software and Apps. We assist with the certification and validation process for Medical Devices and Apps.
While not mentioned in the MDR, the Medical Device Advisor is required in Germany. Manufacturers must appoint and train their MPBs. We offer Medical Device Consultant Training.
The distinction between Medical Devices and Pharmaceuticals can be complex. Differentiation is based on their intended primary effect, with classification depending on this key factor.
Manufacturers must establish and maintain a Quality Management System under ISO 13485 for Medical Devices in the EU market. We ensure compliance and certification.
Regulatory Affairs Consulting for Medical Devices Since 1993, Certified According to ISO 13485
WQS Management Consultants consults manufacturers of medical devices and in vitro diagnostic devices on their path to market access in Europe and the U.S.
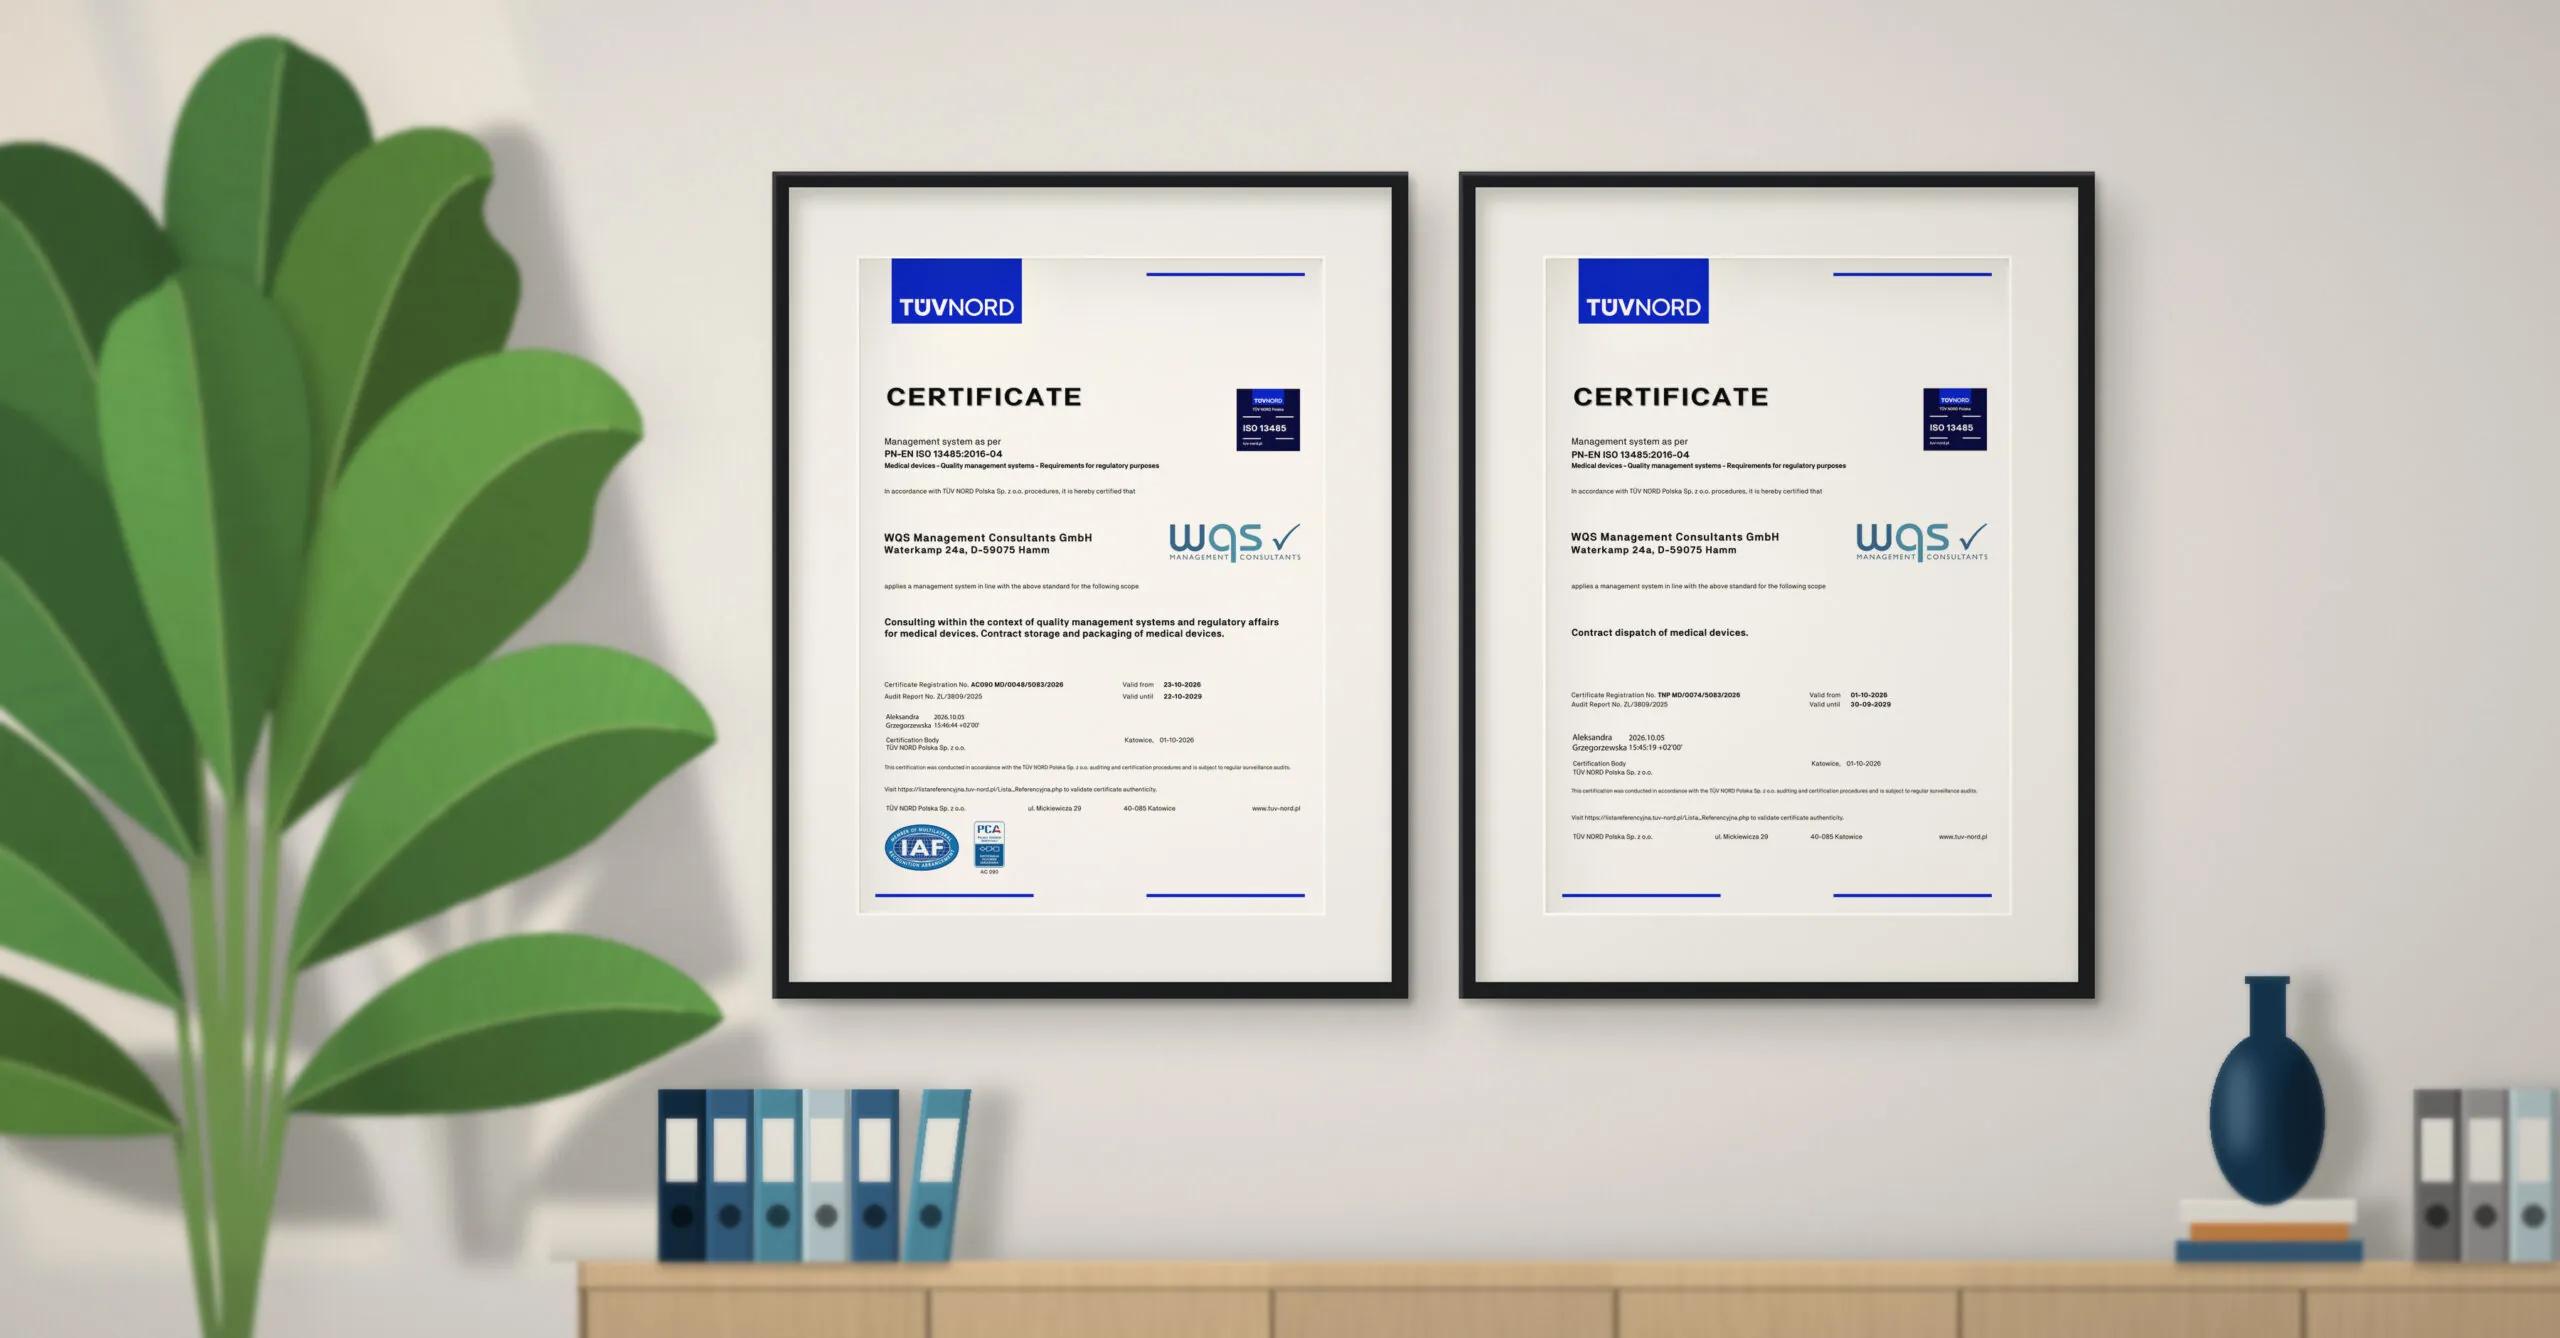
Our own quality management system is certified by TÜV NORD Polska to ISO 13485. The certification covers consulting on quality management and regulatory affairs, as well as the storage and packaging of medical devices on behalf of clients.
WQS Management Consultants advises medical device and IVD manufacturers on regulatory affairs and quality management. FDA, QMSR, ISO 13485, MDR and IVDR. Since 1993, with offices in the United States and Germany.
WQS supports manufacturers with FDA requirements, US Agent representation and the Technical File from an office in St. Petersburg, Florida. The focus is on concrete evidence, not general recommendations.
CE marking requires a full conformity assessment under MDR or IVDR – from classification through technical documentation to the Declaration of Conformity. For US manufacturers, an EU Representative is mandatory.
The technical documentation requires GSPR evidence with product-specific test reports, a clinical evaluation, traceable UDI references and a documented post-market surveillance system.
Risk management under EN ISO 14971 must be demonstrated across the entire product lifecycle as an ongoing process linked to clinical evaluation and post-market surveillance.
For most medical device manufacturers, an ISO 13485 QMS is a precondition for conformity assessment. It covers CAPA processes, document control and audit preparation for Notified Bodies. Under FDA QMSR, ISO 13485 now aligns closely with US requirements.
Depending on the product, evidence on packaging, aging or biocompatibility is required as part of the technical documentation. WQS coordinates the appropriate laboratory tests and assesses the results from a regulatory perspective.
Yes. WQS was founded in Germanyin 1993 and operates from Hamm, North Rhine-Westphalia. That is where the EU regulatory expertise for MDR and IVDR sits.
Manage Cookie Consent
To provide the best experiences, we use technologies like cookies to store and/or access device information. Consenting to these technologies will allow us to process data such as browsing behavior or unique IDs on this site. Not consenting or withdrawing consent, may adversely affect certain features and functions.
Functional
Always active
The technical storage or access is strictly necessary for the legitimate purpose of enabling the use of a specific service explicitly requested by the subscriber or user, or for the sole purpose of carrying out the transmission of a communication over an electronic communications network.
Vorlieben
Die technische Speicherung oder der Zugriff ist für den rechtmäßigen Zweck der Speicherung von Präferenzen erforderlich, die nicht vom Abonnenten oder Benutzer angefordert wurden.
Statistics
Die technische Speicherung oder der Zugriff, der ausschließlich zu statistischen Zwecken erfolgt.The technical storage or access that is used exclusively for anonymous statistical purposes. Without a subpoena, voluntary compliance on the part of your Internet Service Provider, or additional records from a third party, information stored or retrieved for this purpose alone cannot usually be used to identify you.
Marketing
The technical storage or access is required to create user profiles to send advertising, or to track the user on a website or across several websites for similar marketing purposes.